Mental Matters

Health Conditions

Find information on various health conditions affecting people of all ages. This category covers causes, symptoms, diagnosis, and management options for physical illnesses and chronic diseases. Get insights to help you understand different health challenges and make informed decisions about your wellbeing.

Woman in a nightgown sitting thoughtfully on a bed in a softly lit bedroom, suggesting discomfort or concern.

Bacterial Vaginosis Complications

Bacterial Vaginosis Complications – When to See a Doctor While Bacterial Vaginosis is generally not dangerous, it can lead to serious health complications if left untreated — especially for pregnant women and individuals at risk for sexually transmitted infections. Learn more about bacterial vaginosis complications below. Possible complications of untreated or recurrent BV include: You should see a doctor if: BV is not a reflection of cleanliness, and no woman should feel ashamed for seeking care. In South Africa, reproductive health clinics and community-based services can provide confidential, judgement-free treatment. Bacterial Vaginosis Complications Doctors usually don’t treat partners unless the woman experiences repeated reinfections. However, practising safe sex and avoiding irritants can help break the cycle of recurrence. Prompt treatment can clear BV quickly, but maintaining a healthy vaginal microbiome requires understanding and addressing the underlying risk factors. 👉 [Next: Living Well – Preventing BV and Promoting Vaginal Health]

Bacterial Vaginosis Complications Read More »

Black and white photo of a woman partially covering her face with her hands, expressing emotion or introspection.

Living Well with BV

Living Well with BV – Preventing BV and Promoting Vaginal Health Preventing Bacterial Vaginosis involves protecting the natural balance of bacteria in the vagina and avoiding unnecessary disruption. Although not all cases can be avoided, simple lifestyle changes can greatly reduce the likelihood of BV — especially for women who experience frequent recurrences. Read more about living well with BV below. Tips to maintain vaginal health: In South Africa, cultural myths around vaginal hygiene and “tightening” products contribute to harmful practices. Education is critical in dispelling misinformation and promoting healthy, evidence-based habits. Support from clinics, community organisations, and open conversations about women’s health are vital in reducing shame and stigma. Every woman deserves access to factual, respectful information about her body. Living Well with BV Bacterial Vaginosis is common, treatable, and preventable in many cases. With the right knowledge and care, women can take proactive steps to protect their reproductive health and live confidently, comfortably, and well. 👉 [End of Series | Back to Overview]

Living Well with BV Read More »

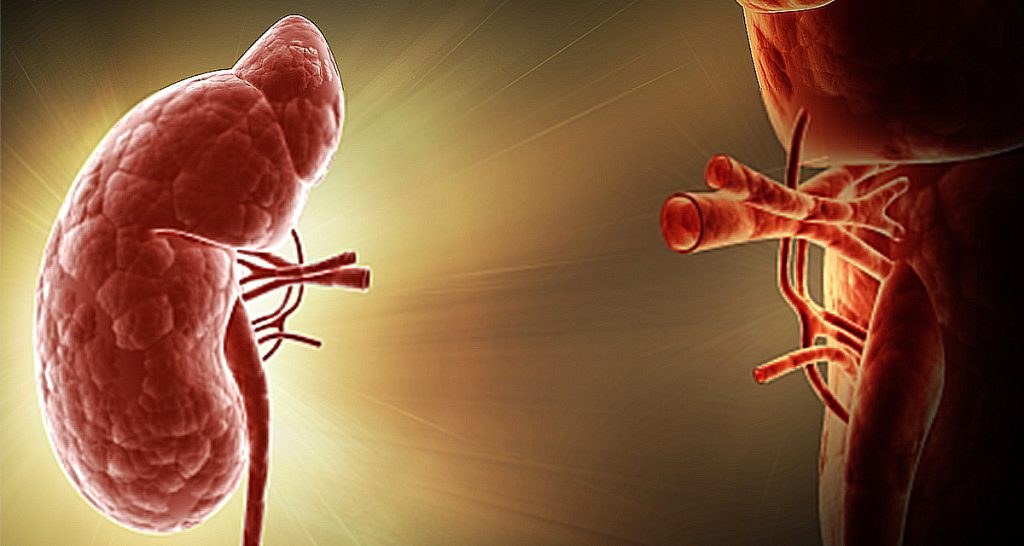
Medical illustration of kidneys highlighting internal structure and arteries, representing ARPKD overview.

Autosomal Recessive Polycystic Kidney Disease (ARPKD) Overview

Autosomal Recessive Polycystic Kidney Disease (ARPKD) Overview Autosomal Recessive Polycystic Kidney Disease (ARPKD) is a rare but serious genetic disorder that primarily affects the kidneys and liver. Unlike the more common autosomal dominant form (ADPKD), ARPKD often presents in infancy or early childhood and follows a recessive inheritance pattern. Meaning a child must inherit two copies of the faulty gene — one from each parent — to develop the condition. ARPKD is characterised by small fluid-filled cysts that develop in the kidney’s collecting ducts. Therefore, causing the kidneys to enlarge and impairing their function. At the same time, it also causes congenital hepatic fibrosis, a progressive liver disease involving bile duct abnormalities and scarring. This dual impact on the kidneys and liver makes ARPKD complex to manage. The severity varies widely — some babies show signs before birth, while others are diagnosed later in childhood. In the most severe cases, symptoms are evident on prenatal scans, and newborns may experience respiratory distress due to underdeveloped lungs caused by the pressure of large kidneys in the womb. Early signs and symptoms include: Autosomal Recessive Polycystic Kidney Disease In South Africa, the early recognition and treatment of ARPKD can be challenging. Especially in rural areas with limited access to neonatal specialists and paediatric nephrologists. In many cases, doctors identify the condition during an ultrasound conducted for another concern. Such as poor growth in utero or abdominal swelling in a newborn. Though ARPKD is life-threatening in its most severe form, many children survive into adolescence and adulthood, especially with early diagnosis. Supportive medical care, and long-term monitoring. Advances in paediatric nephrology and liver management are significantly improving the outlook for affected children. Early Detection Makes a Difference Although ARPKD is a rare condition, recognising the signs early — particularly in newborns and infants — can be life-saving. The combination of kidney and liver involvement means the disease can progress rapidly without appropriate intervention. In South Africa, expanding awareness among healthcare workers. Improving access to prenatal and neonatal screening tools can help identify cases sooner and begin crucial treatment during the most vulnerable stages of life. Living with ARPKD: A Path Forward Despite its complexity, many children with ARPKD go on to lead meaningful lives. With the support of specialist teams, consistent monitoring, and family involvement, long-term outcomes are improving. Continued investment in paediatric nephrology, public education, and outreach programs is essential to bridge the gap between early diagnosis and effective management. Giving more children the opportunity to thrive. 👉 [Next: Causes and Genetics of ARPKD]

Autosomal Recessive Polycystic Kidney Disease (ARPKD) Overview Read More »

Microscopic view of human cells representing genetic and cellular basis of ARPKD.

Causes and Genetics of ARPKD

Causes and Genetics of ARPKD Autosomal Recessive Polycystic Kidney Disease (ARPKD) is caused by mutations in a single gene known as PKHD1. This gene provides instructions for producing a protein called fibrocystin (or polyductin), which is important for the normal development of kidney tubules and bile ducts in the liver. Let’s learn more about the causes and genetics of ARPKD below. Unlike autosomal dominant conditions, ARPKD follows a recessive inheritance pattern. This means that: In most cases, carrier parents are completely healthy and unaware they have a mutated gene. Many only discover their carrier status after having an affected child. For families with a known history of ARPKD, doctors strongly recommend genetic counselling. In South Africa, specialised centres and academic hospitals offer genetic testing, but access is limited in some areas due to costs and the availability of laboratory resources. Many families rely on clinical observation and imaging rather than DNA confirmation. Causes and Genetics of ARPKD The severity of ARPKD doesn’t always correlate with the specific genetic mutation. Two siblings with the same faulty gene may have very different outcomes — one may survive into adulthood, while another may experience complications early in life. It’s important to note that ARPKD is not caused by anything the parents did during pregnancy. It is not related to lifestyle, diet, or environmental exposures. This can be a great comfort to families dealing with feelings of guilt or confusion after diagnosis. Understanding the genetic foundation of ARPKD allows families to make informed decisions about future pregnancies, often with the help of prenatal testing or pre-implantation genetic diagnosis (PGD) if they pursue fertility treatment. Genetics and Family Planning Understanding the recessive inheritance pattern of ARPKD is key to helping families navigate future pregnancies and make informed choices. Genetic counselling offers valuable insight into the risks of recurrence, especially for parents who have already had one affected child. Where available, prenatal testing and pre-implantation genetic diagnosis (PGD) can provide options for early detection or prevention — tools that empower families to prepare with confidence and clarity. Beyond the Genetics: Supporting Families While the condition is genetic, it’s important to reassure families that ARPKD is not the result of anything they did wrong. Guilt is common after a diagnosis, but recognising that the cause lies in unseen genetic mutations can bring some emotional relief. In South Africa, improved access to genetic services and education can help demystify the condition and support early diagnosis — especially in families who may not realise they carry the gene. 👉 [Next: Diagnosis of ARPKD]

Causes and Genetics of ARPKD Read More »

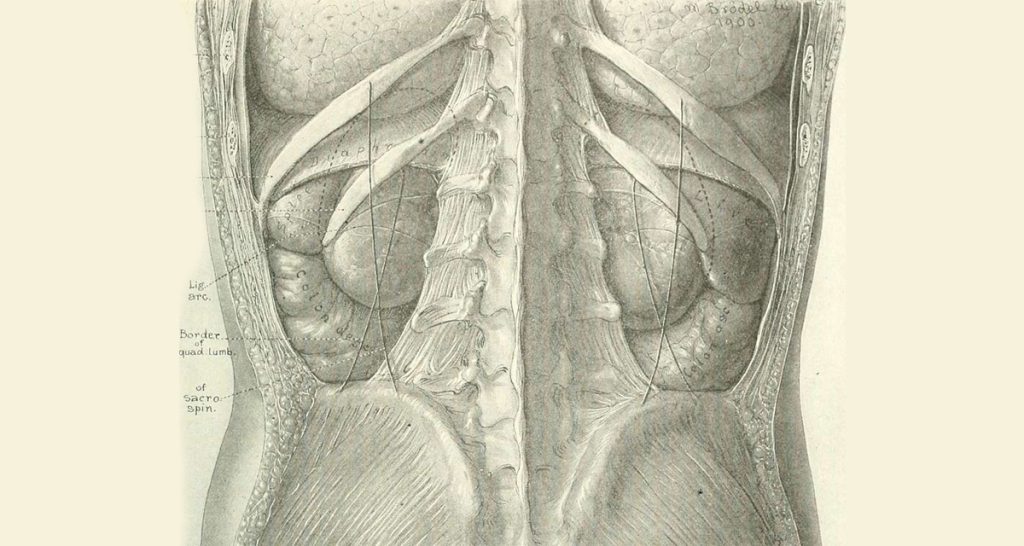
Antique anatomical illustration of the human abdomen showing kidneys, spine, and surrounding tissues.

Diagnosis of ARPKD

Diagnosis of ARPKD Diagnosis of ARPKD typically begins either during pregnancy or shortly after birth, depending on the severity of the condition. Prenatal ultrasounds may detect enlarged, echogenic (bright) kidneys with poor corticomedullary differentiation — a red flag that prompts further investigation. Key diagnostic methods include: Diagnosis of ARPKD In South Africa, limited access to high-resolution ultrasound may delay prenatal diagnosis of ARPKD. After birth, hospitals with paediatric nephrology units—such as those in Cape Town, Johannesburg, or Durban—provide the best care for evaluating and managing suspected cases. Some infants may receive an initial misdiagnosis, especially if born in facilities without paediatric specialists. Once red flags appear, doctors must refer the child promptly to a tertiary hospital. Although ARPKD is a lifelong condition, early diagnosis offers the chance to monitor for complications and begin supportive treatments that can greatly improve long-term outcomes. The Importance of Early and Accurate Diagnosis Early diagnosis of ARPKD lays the groundwork for timely intervention and better long-term management. Identifying the condition prenatally or shortly after birth allows for immediate monitoring of kidney and liver health, as well as blood pressure control — all of which are critical to improving survival and quality of life. Advanced imaging and genetic testing provide clarity for families and medical teams, enabling informed decisions about treatment and care planning. Bridging Gaps in Access and Awareness In South Africa, disparities in access to diagnostic tools remain a challenge, especially in rural or under-resourced areas. Raising awareness among general practitioners, midwives, and sonographers can lead to earlier recognition and referral to specialised centres. With accurate diagnosis and timely support, families affected by ARPKD can begin the journey equipped with knowledge, guidance, and hope for the future. 👉 [Next: Treatment and Ongoing Management of ARPKD]

Diagnosis of ARPKD Read More »

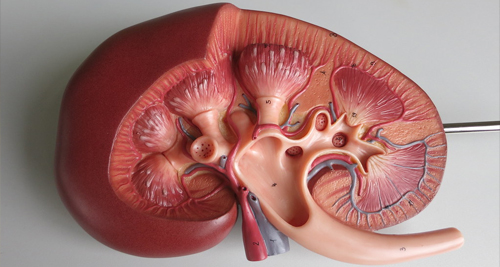
Detailed anatomical model of a human kidney showing internal structures such as calyces, pelvis, and blood vessels.

Treatment and Ongoing Management of ARPKD

Treatment and Ongoing Management of ARPKD There is no cure for ARPKD, but comprehensive medical care can significantly prolong life and improve quality of life, especially when the condition is managed by a multidisciplinary team of nephrologists, hepatologists, dietitians, and therapists. Read more about the treatment and ongoing management of ARPKD below. Treatment depends on the individual’s symptoms and severity, but typically focuses on: Kidney-related treatments include: Liver-related management: Nutrition and growth support are vital. Many children with ARPKD experience failure to thrive, requiring specialised dietary plans or feeding support. In South Africa, access to specialist care varies. Urban centres offer transplant programs and advanced paediatric nephrology services, but rural families may have to travel long distances for dialysis or expert evaluation. Outreach programs and NGO support can help bridge these gaps. Treatment and Ongoing Management of ARPKD Parental education is essential. Caregivers must learn how to monitor fluid intake, recognise signs of infection or high blood pressure, and advocate for their child in healthcare settings. Though the path can be difficult, many children with ARPKD grow into adolescents and even adults — especially with consistent, proactive care and emotional support. Building a Foundation for Better Outcomes Comprehensive and early intervention can make a profound difference in the lives of children with ARPKD. By addressing kidney and liver issues in tandem and supporting nutritional needs, medical teams can help stabilise the condition and reduce the risk of severe complications. Access to specialised care, especially in urban areas of South Africa, plays a crucial role in achieving these outcomes. Supporting Families Every Step of the Way Ongoing education and emotional support empower caregivers to manage the day-to-day demands of ARPKD. From monitoring symptoms to coordinating treatments, families become central partners in care. With the right tools, community resources, and clinical guidance, many children with ARPKD can lead fulfilling lives, gaining the opportunity to grow, learn, and thrive despite the challenges. 👉 [Next: Complications and Prognosis in ARPKD]

Treatment and Ongoing Management of ARPKD Read More »

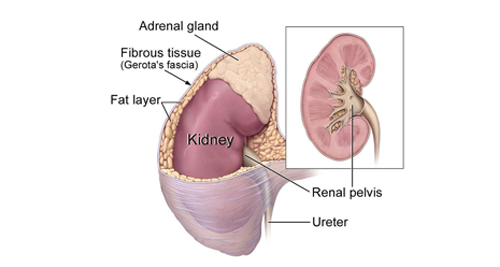
Medical illustration of a human kidney with labelled parts including adrenal gland, renal pelvis, and ureter.

Complications and Prognosis in ARPKD

Complications and Prognosis in ARPKD The course of ARPKD is highly variable. While some infants face life-threatening complications shortly after birth, others live well into adulthood with managed symptoms. Knowing the potential complications and prognosis in arpkd helps families and healthcare providers prepare for and respond to challenges as they arise. Kidney complications: Liver complications: Growth and development: Psychosocial impact: Prognosis depends on several factors: Complications and Prognosis in ARPKD In South Africa, children who reach adolescence with stable organ function may live for decades with good quality of life. Paediatric kidney transplants are successful in major centres, offering long-term relief for some. While the condition is serious, the prognosis is not hopeless. With support, planning, and proactive care, children with ARPKD can live meaningful lives — often filled with resilience, hope, and joy despite the challenges. Navigating Daily Life with Compassion and Structure Living with ARPKD goes far beyond managing medical appointments — it requires a stable routine, emotional resilience, and a deep understanding of a child’s changing needs. Families often become expert caregivers, adapting their homes, schedules, and priorities to meet the complex demands of the condition. Structured daily care, like consistent medication routines and dietary monitoring, helps manage symptoms while giving children the best possible quality of life. Empowerment Through Support and Connection No family should walk this path alone. Support systems — from healthcare providers to community networks and rare disease organisations — play a vital role in sustaining long-term care and emotional wellbeing. In South Africa, these resources may vary by region, but even small connections can offer life-changing reassurance. With the right tools, guidance, and mindset, families can move beyond simply coping, and instead begin building a hopeful, empowered future for their child with ARPKD. 👉 [Next: Living with ARPKD – Family Support and Daily Life] Causes and Genetics of ARPKD Diagnosis of ARPKD Treatment and Ongoing Management of ARPKD Complications and Prognosis of Bile Duct Cancer

Complications and Prognosis in ARPKD Read More »

Artistic representation of healthy and diseased lungs using leaves and twigs on a blue background.

Living with ARPKD – Family Support and Daily Life

Living with ARPKD – Family Support and Daily Life Living with ARPKD requires adaptation, support, and resilience — not just from the child, but from the entire family. While the medical side of the condition is significant, the emotional and social impact is just as important. Daily care includes: Children with ARPKD may miss school for medical appointments or periods of illness. Teachers and caregivers should be educated about the condition and its potential impact on energy levels, focus, and emotional wellbeing. Parental support is critical. Caring for a child with a rare disease can feel overwhelming. Parents often juggle appointments, specialist referrals, diet plans, and medication routines — all while managing their own jobs and relationships. Counselling, support groups, and connecting with other families facing similar journeys can be life-changing. Living with ARPKD In South Africa, resources such as rare disease foundations, hospital social workers, and paediatric outreach clinics can help families access support, education, and funding for treatment where available. Some provincial hospitals partner with NGOs to assist with travel costs, nutritional supplements, or genetic counselling. Hope is central to long-term care. While there is no cure yet, many children live into adulthood, attend school, develop friendships, and experience love and achievement. Planning for the future — including schooling, independence, and mental health — empowers families to look beyond diagnosis. Every child with ARPKD is different. With attentive care, emotional support, and access to evolving treatments, they can thrive in ways that reflect their unique strengths and stories. Navigating Daily Life with Compassion and Structure Living with ARPKD goes far beyond managing medical appointments — it requires a stable routine, emotional resilience, and a deep understanding of a child’s changing needs. Families often become expert caregivers, adapting their homes, schedules, and priorities to meet the complex demands of the condition. Structured daily care, like consistent medication routines and dietary monitoring, helps manage symptoms while giving children the best possible quality of life. Empowerment Through Support and Connection No family should walk this path alone. Support systems — from healthcare providers to community networks and rare disease organisations — play a vital role in sustaining long-term care and emotional wellbeing. In South Africa, these resources may vary by region, but even small connections can offer life-changing reassurance. With the right tools, guidance, and mindset, families can move beyond simply coping, and instead begin building a hopeful, empowered future for their child with ARPKD. 👉 [End of Series | Back to Overview] Autosomal Recessive Polycystic Kidney Disease (ARPKD) Overview Complications and Prognosis in ARPKD Causes and Genetics of ARPKD Diagnosis of ARPKD Treatment and Ongoing Management of ARPKD
Living with ARPKD – Family Support and Daily Life Read More »

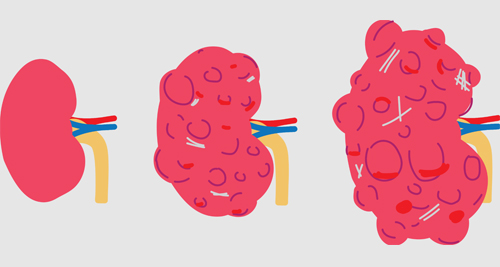
Illustration showing progression of ADPKD from a normal kidney to cyst-filled polycystic kidney.

Autosomal Dominant Polycystic Kidney Disease

Autosomal Dominant Polycystic Kidney Disease (ADPKD) Overview Autosomal Dominant Polycystic Kidney Disease (ADPKD) is a common inherited condition where clusters of fluid-filled sacs — or cysts — develop in the kidneys. These cysts grow over time, often leading to enlarged kidneys, reduced kidney function, and eventually kidney failure. As the name suggests, ADPKD follows an autosomal dominant pattern. This means a child has a 50% chance of inheriting the condition if one parent is affected. 4o mini. The condition affects both kidneys, but one may be more severely impacted than the other. People with ADPKD often remain asymptomatic for decades, with symptoms usually beginning between the ages of 30 and 50. However, early signs can appear in childhood or adolescence, especially in cases where the disease progresses more rapidly. Common symptoms include: In South Africa, many people go undiagnosed for years due to limited access to routine imaging like ultrasounds or CT scans in certain communities. Doctors often discover ADPKD during investigations for hypertension or abdominal pain. Autosomal Dominant Polycystic Kidney Disease ADPKD doesn’t only affect the kidneys. Some people also develop cysts in the liver, aneurysms in the brain, or heart valve abnormalities, making regular monitoring essential. Despite these challenges, many individuals live well for decades with careful medical supervision, lifestyle adjustments, and — when needed — dialysis or transplantation. Managing ADPKD Through Early Diagnosis and Monitoring In conclusion, early diagnosis and regular monitoring of ADPKD are crucial for managing the disease and improving quality of life. With appropriate interventions—such as blood pressure control, pain management, and regular imaging—doctors can slow the progression of kidney damage. Although ADPKD is a lifelong condition with no cure, many individuals with proper care can live well into their later years without reaching kidney failure. The key is staying vigilant and maintaining regular check-ups to catch complications early. The Importance of Genetic Awareness and Family Planning Understanding the genetic nature of ADPKD is vital for affected individuals and their families. Knowing one’s family history and undergoing genetic testing can help inform decisions about family planning and early screening. In South Africa, where access to healthcare may be limited, people must raise awareness about the condition. They must also improve access to genetic counseling and diagnostic tools. With advancements in genetic research and potential new therapies, there is hope for better management and treatment options for future generations. 👉 [Next: Causes and Genetics of ADPKD] Diagnosis of ADPKD Managing and Treating ADPKD Living with ADPKD Autosomal Recessive Polycystic Kidney Disease (ARPKD) Overview

Autosomal Dominant Polycystic Kidney Disease Read More »

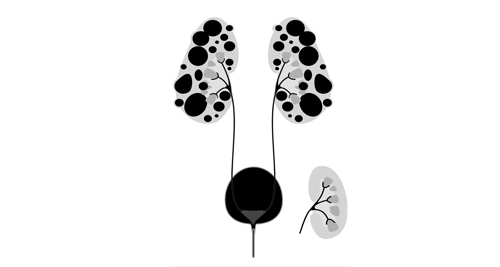
Illustration of kidneys with multiple cysts representing Autosomal Dominant Polycystic Kidney Disease (ADPKD) and genetic cause.

Causes and Genetics of ADPKD

Causes and Genetics of ADPKD ADPKD is a genetic condition, passed from one generation to the next through mutations in specific genes. Learn more about the Causes and Genetics of ADPKD below. The two primary genes involved are: Mutations in PKD1 generally result in more severe disease with earlier onset of symptoms and faster progression to kidney failure. PKD2 mutations tend to be milder, with later onset and slower progression. Both forms follow the autosomal dominant inheritance pattern, meaning that only one faulty copy of the gene is needed to cause the disease. In rare cases, ADPKD can also occur as a spontaneous mutation, meaning there is no known family history. Unlike some genetic disorders that skip generations, ADPKD is usually passed down directly from one generation to the next. If one parent carries the mutation, each child has a 50% chance of inheriting the disorder. A person with the disease can also pass it to their children even if their symptoms are mild. Causes and Genetics of ADPKD Family history is a major clue in the diagnostic process. If a close relative has been diagnosed, it’s often recommended that family members seek genetic counselling and, in some cases, undergo genetic testing. However, this isn’t always available in South Africa’s public sector, and imaging studies like ultrasounds or CT scans are still the primary diagnostic tools. Understanding the genetic basis of ADPKD can help patients and families make informed decisions about family planning, screening, and long-term care. It also paves the way for potential targeted therapies, some of which are already being trialled internationally. Understanding the Genetic Foundation of ADPKD ADPKD is a hereditary condition that follows an autosomal dominant inheritance pattern, meaning that individuals with one faulty gene from either parent are at risk. The two primary genes responsible for this condition, PKD1 and PKD2, lead to different severities and ages of onset. While most cases are inherited, spontaneous mutations can occur, so it’s important to consider family history during diagnosis. Genetic testing, while invaluable, may not always be accessible in some regions, which makes diagnostic imaging such as ultrasounds and CT scans vital tools in early detection. Importance of Genetic Knowledge for ADPKD Management Understanding the genetic causes of ADPKD enables better management and care strategies for patients. Knowing one’s genetic predisposition helps in making informed decisions about family planning and potential screening for future generations. Furthermore, genetic knowledge can pave the way for developing targeted therapies, which hold the promise of slowing down the disease progression. In South Africa, patients may have limited access to genetic counselling and advanced testing. However, ongoing advancements in understanding the genetic roots of ADPKD continue to shape the future of treatment and care. 👉 [Next: Diagnosis of ADPKD] Living with ADPKD Managing and Treating ADPKD Complications and Long-Term Outlook of ADPKD Autosomal Dominant Polycystic Kidney Disease

Causes and Genetics of ADPKD Read More »

Scroll to Top