Mental Matters

Idiopathic Pulmonary Fibrosis

Medical professional holding stethoscope next to text reading Idiopathic Pulmonary Fibrosis

Idiopathic Pulmonary Fibrosis

Overview of Idiopathic Pulmonary Fibrosis Idiopathic pulmonary fibrosis is a rare, long-term lung condition in which the tissue deep within the lungs becomes thickened, stiff, and scarred over time. What Is Idiopathic Pulmonary Fibrosis? The word “idiopathic” means the cause is unknown. “Pulmonary fibrosis” means scarring in the lungs. This scar tissue harms lung structure. It makes it hard for oxygen to pass from the air sacs into the blood. As a result, people with idiopathic pulmonary fibrosis often have growing shortness of breath, a dry cough that won’t go away, and worsening tiredness. How Idiopathic Pulmonary Fibrosis Fits Into Lung Diseases Idiopathic pulmonary fibrosis belongs to a group called interstitial lung diseases (ILDs). These diseases cause inflammation or scarring in the lung’s support tissue. Some ILDs have known causes, like exposure to harmful dust, autoimmune diseases, or medicines. But IPF happens without a known trigger. Doctors usually diagnose it after ruling out other causes. This process can take time. Who Is Affected? IPF mostly affects people over 50. It happens more in men than women. It is more common in people who have smoked or breathed in dust like wood or metal dust for a long time. Though rare, IPF has gained more attention due to better diagnosis and research. Still, it remains serious. Most people live 3 to 5 years after diagnosis. How IPF Progresses IPF changes unpredictably. Some people worsen slowly over years. Others may suddenly lose lung function fast. Sometimes, sudden attacks called acute exacerbations happen. These can be life-threatening. Because of this, regular check-ups with lung specialists are very important. Early Symptoms and Signs The early symptoms are mild and can look like other lung problems such as asthma or bronchitis. Early signs include breathlessness during exercise and a dry cough that does not bring up mucus. Over time, everyday tasks become harder. Some people develop finger or toe clubbing—a swelling caused by low oxygen over time. Diagnosing IPF Doctors use many tools to diagnose IPF. These include clinical exams, high-resolution CT scans, lung function tests, and sometimes lung biopsies. It is important to tell IPF apart from other lung diseases because treatment and outlook differ. Today, teams of lung doctors, radiologists, and pathologists work together to make a firm diagnosis and plan care. Treatment Options There is no cure for idiopathic pulmonary fibrosis. But new drugs like pirfenidone and nintedanib can slow down lung damage. These medicines do not fix the scars but help slow worsening. Oxygen therapy, lung rehab, and lifestyle changes also help manage symptoms. In severe cases, lung transplant may be an option. Living with IPF Living with IPF is tough both physically and emotionally. It limits movement and independence. It may cause anxiety, depression, and social isolation. Support from family, patient groups, and mental health professionals is key. Learning about IPF helps patients take charge of their care and make informed choices. What’s Next? In the following sections, we will cover the causes, symptoms, diagnosis, treatment, complications, and outlook of idiopathic pulmonary fibrosis. This guide aims to help patients, caregivers, and healthcare workers understand and manage this complex disease. [Next: Causes of Idiopathic Pulmonary Fibrosis →]
Idiopathic Pulmonary Fibrosis Read More »

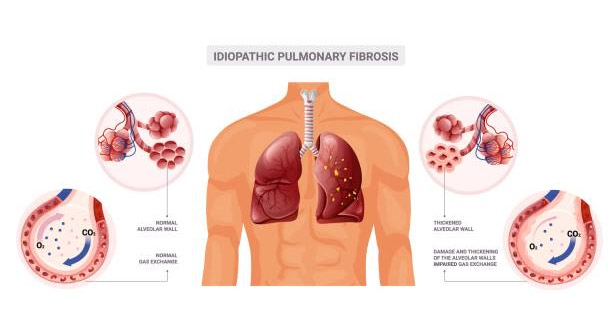
Medical illustration showing lungs affected by idiopathic pulmonary fibrosis and alveolar damage

Causes of Idiopathic Pulmonary Fibrosis

Causes of idiopathic pulmonary fibrosis remain largely unknown, which is reflected in the term “idiopathic”—meaning no identifiable origin. Why Causes Are Hard to Find Many factors may cause IPF, but no single cause explains it all. Scientists think it comes from a mix of genetics, environment, immune problems, and ageing. Knowing the causes of idiopathic pulmonary fibrosis helps with early detection and better treatments. How the Disease Starts Researchers believe IPF begins with repeated tiny injuries to the lungs. Instead of healing well, the lungs repair wrongly. Special cells called fibroblasts make too much scar tissue. This scarring stiffens the lungs and makes breathing hard. But what triggers this faulty healing is still unknown. Role of Genetics Genes give clues to why some people get IPF. Mutations in genes like TERT, TERC, and MUC5B raise the risk. These gene changes can shorten lung cell life or cause too much mucus. People with family history of lung disease often share these mutations. Environmental and Workplace Risks Long-term exposure to dust, metal, farming materials, silica, and chemicals may increase risk. Workers in carpentry, farming, or construction often have higher chances of IPF. These irritants may cause ongoing lung inflammation and damage in people who are already vulnerable. Smoking as a Risk Factor Many IPF patients have a history of smoking, though not all. Smoking harms lung cells and slows healing. It may also speed up IPF and reduce how well treatments work. Tobacco is a major risk factor but not the only cause. Immune System Involvement The immune system may react wrongly in IPF. Instead of fixing damage, it may cause more scarring. Autoimmune diseases like rheumatoid arthritis can cause similar lung problems. But in true IPF, the immune trigger is unclear. Viruses and hidden inflammation may also play a role. Age and IPF IPF usually develops after age 50 and gets more common with age. Older lungs may not repair damage well. They also carry genetic changes from years of exposure. Ageing itself may make lungs more likely to scar after injury. Other Possible Causes Gastroesophageal reflux disease (GERD) may add to lung damage. Acid from the stomach might leak into lungs, harming tissue. Treating reflux could reduce flare-ups. Some viruses, like Epstein-Barr and hepatitis C, are found more often in IPF lungs. Their exact role is still being studied. Uncertainty Remains Most of the time, no clear cause is found. This makes diagnosis and treatment harder. Research continues to explore all possible factors behind IPF. Summary of Causes of Idiopathic Pulmonary Fibrosis The causes of idiopathic pulmonary fibrosis involve many factors. Genes, environment, immune problems, and ageing all play parts. Understanding these helps detect IPF early and develop better treatments. Future research may uncover new ways to prevent and manage this disease. [Next: Symptoms of Idiopathic Pulmonary Fibrosis →]
Causes of Idiopathic Pulmonary Fibrosis Read More »

Elderly man coughing and holding chest, showing signs of idiopathic pulmonary fibrosis

Symptoms of Idiopathic Pulmonary Fibrosis

Symptoms of idiopathic pulmonary fibrosis often develop gradually and can be subtle in the early stages, which makes early recognition and diagnosis particularly challenging. Why Symptoms Are Missed Early Many people think these symptoms are due to age, poor fitness, or common lung issues like asthma or bronchitis. As the disease moves forward, the signs grow stronger and more disruptive. Knowing the main symptoms of idiopathic pulmonary fibrosis is vital for early diagnosis and better care. Understanding these signs helps both patients and doctors act quickly. Shortness of Breath The most common early sign is breathlessness, also called dyspnoea. At first, this happens only during activities like climbing stairs or walking fast. Later, it becomes constant, even at rest. This occurs because lung tissue turns stiff and cannot expand well. Oxygen flow drops, and simple tasks become tiring. Many patients need frequent breaks just to finish daily activities. Persistent Dry Cough Another key symptom of idiopathic pulmonary fibrosis is a dry cough that does not produce mucus. This cough often stays for months and does not respond to usual cough medicines. It can disturb sleep, interrupt talking, and even get worse in cold air or while laughing. Unlike the wet cough in infections, this cough feels harsh and hard to manage. It can also cause social stress and emotional strain. Fatigue and Low Energy Tiredness is another major symptom. The body works harder to get oxygen, which strains the heart and breathing muscles. Over time, people feel drained even after rest. Low oxygen delivery adds to this fatigue, making daily life harder. This can lead to less work activity and social withdrawal. Fatigue is often underestimated but impacts quality of life as much as breathing problems. Visible Physical Changes As IPF worsens, some people develop finger clubbing. The fingertips enlarge, and nails curve around them. This sign happens due to long-term oxygen shortage. While it does not hurt, it signals serious lung problems. Another late sign is cyanosis, where lips, nails, or skin turn bluish due to very low oxygen. People may also breathe faster and use neck or chest muscles to help with breathing. Other Symptoms and Emotional Impact Some patients lose weight without trying or feel chest discomfort. Others simply feel unwell most of the time. The mental impact is also strong. Breathlessness and tiredness can lead to anxiety and depression. Worry about sudden flare-ups makes stress worse. Good IPF care should include both physical and emotional support. Variations and Sudden Worsening Symptoms of idiopathic pulmonary fibrosis do not progress the same way in everyone. Some decline slowly; others stay stable and then worsen quickly. Acute flare-ups can happen without warning and often require hospital care. These sudden drops in lung function are dangerous and may need high-dose steroids or breathing machines. Why Misdiagnosis Happens Early IPF symptoms look like those of COPD, asthma, or heart failure. This can cause delays in diagnosis. Anyone over 50 with lasting breathlessness, cough, or past exposure to smoking or pollutants should get lung tests and imaging. Final Thoughts on Symptoms of Idiopathic Pulmonary Fibrosis Knowing the symptoms of idiopathic pulmonary fibrosis early helps start treatment sooner. Breathlessness, dry cough, fatigue, finger clubbing, and blue lips are all warning signs. The earlier these symptoms are found, the better the chance to slow the disease and improve life. [Next: Diagnosis of Idiopathic Pulmonary Fibrosis →]
Symptoms of Idiopathic Pulmonary Fibrosis Read More »

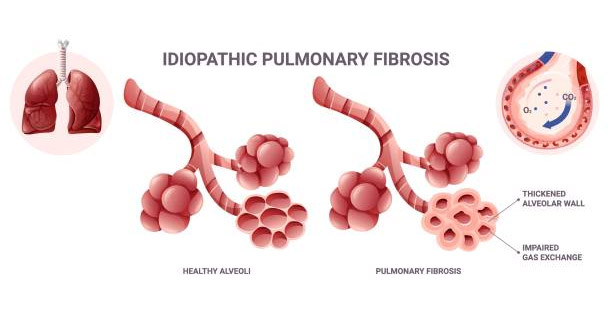
Medical illustration comparing healthy alveoli and fibrotic lungs in idiopathic pulmonary fibrosis

Diagnosis of Idiopathic Pulmonary Fibrosis

Diagnosis of idiopathic pulmonary fibrosis is often complex and requires a multifaceted approach to rule out other conditions that may mimic its symptoms. Why Diagnosis Is Difficult Idiopathic pulmonary fibrosis (IPF) looks similar to many other lung diseases, especially interstitial lung disease (ILD). This makes accurate diagnosis challenging. Early symptoms such as breathlessness and tiredness are often mild. Many people ignore them or think they are due to aging. Still, an early and correct diagnosis of idiopathic pulmonary fibrosis is critical for starting treatment and slowing the disease. Medical History and Physical Exam The first step in diagnosing IPF starts with a detailed medical history and physical check-up. Doctors ask about long-term symptoms like dry cough and shortness of breath. They also ask about exposure to dust, smoke, or chemicals, past smoking habits, family history, and any autoimmune diseases. These details help rule out other lung problems that may need different treatment. During the exam, a doctor may hear “Velcro-like” crackles in the lungs with a stethoscope. These soft sounds happen when a person breathes in and often suggest IPF. Another sign is finger clubbing, where fingertips become round and swollen because of low oxygen. These clues do not confirm IPF, but they make doctors more suspicious when combined with symptoms and history. Imaging Tests To confirm IPF, doctors usually order a high-resolution CT scan (HRCT). This special scan shows detailed images of lung tissue. In IPF, HRCT often shows a usual interstitial pneumonia (UIP) pattern. This includes honeycombing, thickened tissue, and airway changes. When this pattern is clear, doctors can often confirm IPF without other invasive tests. When Imaging Is Not Enough If the scan does not clearly show UIP, doctors may suggest a lung biopsy. This can be done by surgery or with a less invasive cryobiopsy. A small piece of lung tissue is studied under a microscope to confirm IPF. But biopsies carry risks, especially in older patients or those with poor lung function. Doctors weigh these risks before deciding. Pulmonary Function Tests Another key step is lung function testing. These tests check how well the lungs move air and transfer oxygen. IPF patients often have stiff lungs that cannot expand well. This shows as low forced vital capacity (FVC) and poor oxygen transfer (DLCO). Repeat tests over time help track disease progress. Blood Tests and Other Checks Blood tests rule out other diseases that look like IPF, such as autoimmune disorders. They check for antibodies and inflammation markers. But no blood test can confirm IPF yet. Researchers are studying new markers, but these are not in routine use now. Role of Multidisciplinary Teams Because IPF is hard to diagnose, many patients are referred to a team of specialists. This team often includes lung doctors, radiologists, pathologists, and sometimes rheumatologists. They review scans, lab tests, and symptoms together. This combined approach is now the best way to confirm IPF and plan the right treatment. Bronchoalveolar Lavage (BAL) Sometimes doctors use BAL to collect lung fluid for testing. It cannot confirm IPF on its own, but it helps rule out infections or other lung diseases. It is most useful in younger patients or when the case is unclear. Importance of Early Diagnosis Getting diagnosed early makes a big difference in treatment and quality of life. Sadly, many patients face long delays. Symptoms are often blamed on age or asthma. Lack of specialist access also slows diagnosis. Raising awareness among doctors and patients can help reduce these delays. Final Thoughts on Diagnosis of Idiopathic Pulmonary Fibrosis The diagnosis of idiopathic pulmonary fibrosis needs a full review of symptoms, scans, lung tests, and sometimes biopsies. A specialist team often gives the most accurate answer. The sooner IPF is found, the sooner treatment can begin to manage symptoms and improve life quality. [Next: Treatment of Idiopathic Pulmonary Fibrosis →]
Diagnosis of Idiopathic Pulmonary Fibrosis Read More »

Antibiotic capsules and medicine bottle labelled for idiopathic pulmonary fibrosis treatment

Treatment of Idiopathic Pulmonary Fibrosis

Treatment of idiopathic pulmonary fibrosis focuses on slowing disease progression, managing symptoms, improving quality of life, and preventing complications. Overview of Current Treatment There is no cure for idiopathic pulmonary fibrosis (IPF). The disease usually worsens with time. However, new treatments now help slow lung damage and improve life for many patients. Treatment of idiopathic pulmonary fibrosis often needs a team approach. This includes lung specialists, therapists, pharmacists, and care experts. Antifibrotic Medications The main treatment for idiopathic pulmonary fibrosis includes antifibrotic medicines. These medicines help slow lung scarring. Two key drugs are pirfenidone and nintedanib. Pirfenidone lowers lung inflammation and slows scarring. Nintedanib blocks several pathways that lead to fibrosis. Both medicines can slow disease progression but cannot reverse lung damage or cure IPF. Still, they give patients more time and better stability. Managing Side Effects These medicines can cause side effects. Pirfenidone may cause nausea, tiredness, and skin sensitivity to sunlight. Nintedanib often leads to diarrhoea and high liver enzymes. Doctors monitor patients closely and adjust doses when needed. Regular check-ups help track lung function, review side effects, and improve medicine tolerance. Pulmonary Rehabilitation Supportive care is also very important. Pulmonary rehabilitation is a structured program with breathing exercises, physical activity, diet tips, and mental health support. It improves stamina, lowers breathlessness, and boosts quality of life. Joining a rehab program early helps patients stay active and independent longer. Oxygen Therapy for IPF As the disease gets worse, many patients need extra oxygen. Oxygen therapy helps maintain blood oxygen levels during sleep and activity. Portable oxygen systems allow mobility and daily activities. Long-term oxygen also reduces heart strain and prevents complications such as pulmonary hypertension. Doctors decide on oxygen needs using simple tests like pulse oximetry. Infection Prevention Preventing infections is key in the treatment of idiopathic pulmonary fibrosis. Infections can cause sudden worsening of IPF. Patients should get flu and pneumonia vaccines every year. Quick treatment of chest infections and avoiding crowded places helps reduce risk. Good hand hygiene and quitting smoking are vital steps for lung health. Managing Other Health Issues Other conditions often come with IPF, like GERD, sleep apnea, heart disease, and pulmonary hypertension. GERD may worsen lung scarring. Doctors may suggest medicines such as proton pump inhibitors or anti-reflux surgery. Treating these conditions improves health and slows IPF progression. Role of Palliative Care Palliative care is part of IPF treatment, even early on. It focuses on comfort, reducing symptoms, and supporting emotional health. Palliative care offers medicine for breathlessness, counseling, and family support. It does not mean stopping treatment but working alongside it for a better life. Lung Transplantation Lung transplantation is the only option that may add years of life for some patients. It is for those with fast disease worsening or severe oxygen needs. Doctors check age, overall health, and response to treatment before recommending it. While it can improve life, the surgery has big risks, including organ rejection and medicine side effects. Future Treatments and Research Research for idiopathic pulmonary fibrosis is growing fast. New drugs that target fibrosis pathways, immune changes, and cell repair are in trials. Patients can join clinical trials to try new treatments and help improve future care. Education and Self-Management Patient education matters a lot in IPF care. People should understand their illness, medicine schedule, and early warning signs. Knowing when to call a doctor can save lives. Healthy habits, like eating well, saving energy, and avoiding very high places or cold air, also help daily living. Final Thoughts on Treatment of Idiopathic Pulmonary Fibrosis The treatment of idiopathic pulmonary fibrosis needs a full, team-based plan. Medicines, rehab, oxygen, infection control, and emotional care all play a part. For advanced cases, transplant and palliative care help improve comfort and survival. IPF is still a tough disease, but early care and ongoing support can make a real difference. [Next: Complications of Idiopathic Pulmonary Fibrosis →]
Treatment of Idiopathic Pulmonary Fibrosis Read More »

Illustration of a patient on oxygen therapy for complications of idiopathic pulmonary fibrosis

Complications of Idiopathic Pulmonary Fibrosis

Complications of idiopathic pulmonary fibrosis often arise as the disease progresses, significantly impacting both prognosis and quality of life. Why Complications Develop While idiopathic pulmonary fibrosis (IPF) mainly causes lung scarring and breathlessness, its effects spread far. They can even lead to life-threatening problems if ignored. IPF is a long-term and worsening disease, so complications build slowly, though sudden flare-ups can happen. Understanding the complications of idiopathic pulmonary fibrosis helps both doctors and patients manage risks and plan treatment. Acute Exacerbations One of the most serious complications of idiopathic pulmonary fibrosis is an acute flare-up. These sudden episodes cause fast worsening of breathing and can be deadly. People often feel more short of breath and have very low oxygen levels. Scans like a CT often show new lung changes. These attacks may look like infections, but many happen without a clear reason. Most patients need hospital care with strong steroid medicines and oxygen. Sadly, these episodes have poor outcomes and remain a leading cause of death in IPF. Pulmonary Hypertension Another major complication is pulmonary hypertension. This means high blood pressure in the lung arteries. As lung scarring grows and oxygen drops, small lung blood vessels tighten or break down. This raises pressure and strains the heart, often causing right-sided heart failure (cor pulmonale). Treating pulmonary hypertension is hard and usually means advanced disease. Common signs include swelling in the legs, chest pain, and severe tiredness. Doctors use oxygen and certain drugs to ease the heart and lung stress, but results are often limited because of the damage. Gastro-Oesophageal Reflux (GERD) GERD is another overlooked complication of idiopathic pulmonary fibrosis. Many patients suffer from acid reflux. Research shows stomach acid may reach the lungs and make scarring worse. GERD can be silent, so people might not notice symptoms. Still, acid can harm the lungs over time. Treating GERD with medicines like proton pump inhibitors or, in some cases, surgery (such as Nissen fundoplication) can help lower flare-ups and improve comfort. This makes GERD a fixable risk in IPF care. Infections and Their Risks Infections are a big worry for people with idiopathic pulmonary fibrosis. Because their lungs work poorly and clear mucus slowly, they face higher infection risks. Pneumonia is common. Those taking steroids or immune-suppressing drugs have extra risk of viral or fungal infections. Infections can trigger sudden decline and worsen breathing. Preventive steps like flu and pneumonia vaccines, quick treatment of chest symptoms, and good hygiene are key. Doctors must act fast since symptoms can appear mild at first in IPF. Lung Cancer in IPF Lung cancer is also common in patients with idiopathic pulmonary fibrosis, especially smokers. Ongoing lung injury and scarring may cause cancer cells to grow. Detecting cancer in IPF patients is tough because the symptoms—like cough or weight loss—are similar to IPF. Biopsies are risky for fragile lungs, and treatment options such as chemo or surgery can worsen fibrosis. So, treatment must be very careful and tailored to the patient. Mental and Emotional Impact Idiopathic pulmonary fibrosis does not only harm the lungs. It deeply affects mental health. Many patients feel depressed or anxious due to the long-term illness, loss of independence, and fear of the future. Fatigue and breathlessness can stop social life and make daily tasks very hard. Support through therapy, support groups, or medicine can greatly improve life. Mental health care should always be part of IPF treatment. End-Stage Respiratory Failure In the last stage of IPF, breathing failure becomes the biggest threat. The lungs can no longer give enough oxygen, even with high-flow oxygen therapy. Patients feel breathless all the time, extremely tired, and unable to move much. Palliative care becomes vital, focusing on comfort and emotional support. Lung transplant may help some people, but most are not eligible because of age or other health problems. Spotting this stage early helps families plan care and prepare for end-of-life support. Caregiver Strain Idiopathic pulmonary fibrosis also affects families. As patients need more care, loved ones often face burnout, stress, and money problems. Education, counseling, and rest options for caregivers are very important. Involving family in care planning helps them feel ready and supported. Blood Clots and Mobility Issues Sometimes, IPF patients develop blood clots like pulmonary embolism or deep vein thrombosis (DVT). This happens when people move less and have ongoing inflammation. To lower this risk, patients should stay as active as possible, use compression stockings, and take blood thinners if needed. These clots are dangerous because IPF patients already struggle to breathe. Final Thoughts on Complications of Idiopathic Pulmonary Fibrosis The complications of idiopathic pulmonary fibrosis affect both body and mind. Acute flare-ups, pulmonary hypertension, infections, GERD, lung cancer, and end-stage failure all threaten health and life. Mental distress and caregiver stress add more challenges. By spotting and treating these risks early, doctors can improve life quality and help patients and families manage this tough disease. [Next: Outlook for Idiopathic Pulmonary Fibrosis →]

Complications of Idiopathic Pulmonary Fibrosis Read More »

Doctor discussing prognosis with a smiling patient recovering from idiopathic pulmonary fibrosis

Outlook for Idiopathic Pulmonary Fibrosis

The outlook for idiopathic pulmonary fibrosis (IPF) remains guarded because the disease is progressive and currently incurable. However, new treatments and better disease awareness have improved symptom control, quality of life, and in some cases, survival. Each patient’s outcome depends on factors such as age, overall health, speed of disease progression, and access to specialist care. Understanding the outlook for idiopathic pulmonary fibrosis helps patients and families plan ahead and make informed decisions. Life Expectancy and Disease Progression On average, life expectancy after an IPF diagnosis is three to five years, but this number varies widely. Some people live much longer, especially with early diagnosis and treatment, while others experience rapid decline. Prognosis is unpredictable and can change suddenly. Doctors often use the GAP index (Gender, Age, Physiology) to estimate risk, but even this model cannot predict every case. Treatments That Improve Outlook The biggest advance in improving outlook for IPF is the use of antifibrotic drugs—pirfenidone and nintedanib. These medicines do not cure the disease but slow lung function decline, reduce hospitalisations, and may extend survival. Starting treatment early, before severe lung damage occurs, offers the most benefit. Other important treatments include: Patients who stay active, maintain good nutrition, and receive regular specialist care often report better physical and emotional well-being. Complications and Their Impact Complications such as acute exacerbations, pulmonary hypertension, and lung cancer can shorten survival and increase hospital stays. Preventing these events—through infection control, healthy lifestyle habits, and regular monitoring—helps improve prognosis. Role of Lung Transplantation For eligible patients, a lung transplant offers the best chance for long-term survival and better quality of life. Transplant recipients can live many years beyond the typical course of IPF. However, not everyone qualifies due to age, other medical conditions, or late diagnosis. Early referral to a transplant centre is essential. Mental Health and Emotional Well-being The emotional toll of IPF is significant. Depression, anxiety, and social withdrawal are common and can reduce treatment adherence. Counselling, peer support groups, and mindfulness can help patients cope and improve their outlook on life. Planning for the Future Advance care planning is crucial. Discussing treatment goals, palliative care options, and end-of-life wishes early provides patients and families with peace of mind. This proactive approach ensures dignity, control, and emotional preparedness as the disease progresses. Hope for the Future Research continues to explore new drugs targeting fibrosis and immune pathways. Genetic studies and biomarkers may soon allow personalised treatment. Clinical trials are testing therapies aimed at slowing or even reversing lung damage. Final Thoughts Although the outlook for idiopathic pulmonary fibrosis remains serious, it is improving. With early diagnosis, antifibrotic therapy, proactive complication management, and strong emotional support, many people live longer, more fulfilling lives. The future of IPF care looks promising as science moves closer to more effective treatments and, possibly, a cure. [Next: Back to Overview →]

Outlook for Idiopathic Pulmonary Fibrosis Read More »

Scroll to Top